September 14th, 2026
By Kloie Bautz
Bringing a medical device from early development toward clinical use involves many different kinds of testing.
Some studies answer whether a technology works. Others help engineers refine a design, verify performance, or better understand potential risks. Then there are studies intended to generate safety evidence that may ultimately be submitted to the FDA.
That is where Good Laboratory Practice, or GLP, can come into the picture.
So what makes a study GLP? How is it different from the other testing performed during medical device development? And why might the FDA expect one?
This primer provides a starting point for understanding where GLP fits into the larger medical device development process.
GLP stands for Good Laboratory Practice.
In the United States, the FDA’s GLP requirements are established in 21 CFR Part 58, Good Laboratory Practice for Nonclinical Laboratory Studies.
GLP applies to certain nonclinical laboratory studies that evaluate safety. These studies are intended to support research or marketing applications for FDA-regulated products, including medical devices.
The purpose is to help ensure the quality and integrity of the safety data submitted to the FDA.
Importantly, GLP is not a particular experiment or test method. There is no single test called a “GLP test.”
Instead, GLP establishes a framework for how an applicable study is planned, conducted, monitored, documented, reported, and archived.
Requirements address areas such as:
In practical terms, GLP creates a documented and traceable system around a study. This helps regulators understand both the results and how those results were generated.
Medical device development includes many kinds of nonclinical testing, and those studies can serve very different purposes.
Early in development, a team may be asking questions such as:
These are often exploratory or feasibility questions. The goal is to learn and improve the technology, so flexibility can be important while the design is still changing.
Later, engineering and verification testing may answer questions such as:
This may include bench testing, software testing, mechanical testing, electrical testing, imaging performance testing, or other device-specific evaluations.
These studies can be highly controlled and well documented without necessarily being GLP studies.
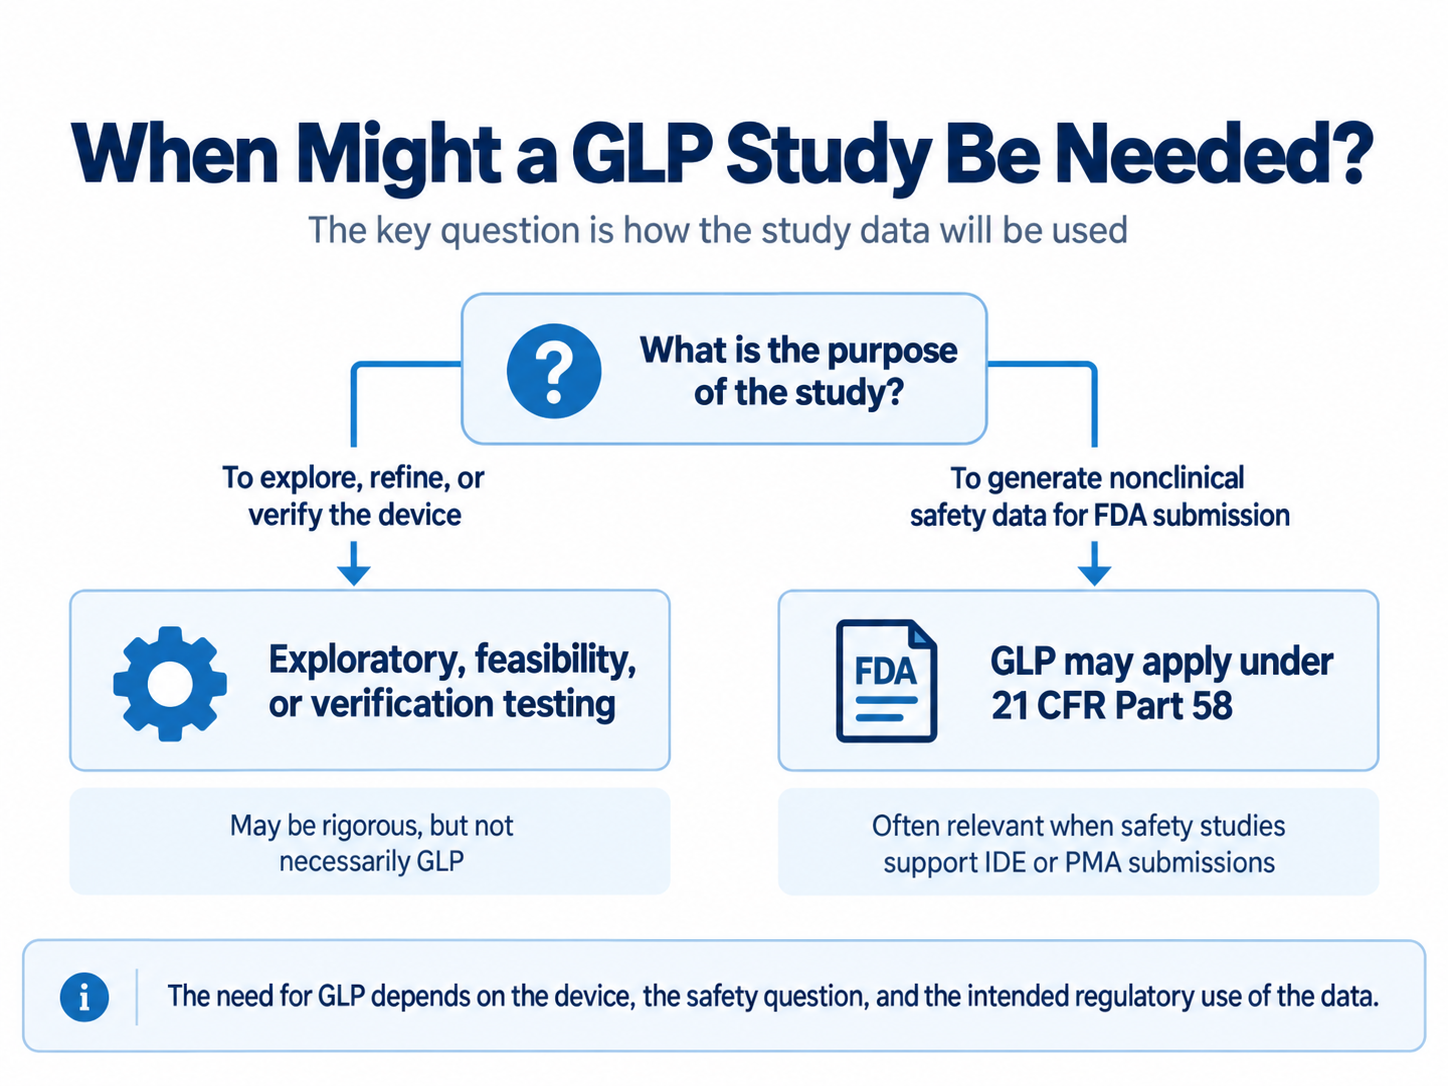
GLP becomes particularly relevant when a nonclinical study is intended to generate safety data to support an FDA research or marketing application.
The difference is not that one type of study represents “good science” and another does not.
Rather, they serve different purposes.
A non-GLP study may be exactly what is needed during early development.
For example, engineers may intentionally test several prototype configurations, modify parameters between experiments, or change procedures as they learn more about a technology.
That flexibility is often valuable during research and development.
A study intended to generate formal regulatory safety evidence has a different purpose. Its results may need to stand on their own as part of the evidence FDA evaluates.
A useful way to think about the difference is:
Two studies can investigate a similar scientific question but differ substantially in how they are conducted.
A GLP study adds formal controls throughout the study lifecycle.
Before testing begins, the study is conducted according to a written protocol defining its objectives and planned methods.
The protocol may describe the test and control articles, experimental design, procedures, measurements, endpoints, and other study-specific requirements.
GLP establishes responsibilities for individuals involved in the study.
For example, a study director has overall responsibility for the conduct of the study and serves as its central point of control.
Management, technical personnel, and quality assurance personnel also have defined roles.
Equipment must be appropriately maintained and, where necessary, inspected and calibrated.
Routine laboratory procedures are controlled through written standard operating procedures, or SOPs.
Observations and measurements must be recorded in a way that preserves what occurred during the study.
Changes, corrections, and deviations are also documented.
The goal is for the study record to show what happened rather than relying on someone’s memory after the fact.
GLP also requires a Quality Assurance Unit, or QAU, that operates independently from the personnel conducting the study.
Quality assurance personnel inspect the study and help verify that it is being performed according to the protocol, SOPs, and applicable GLP requirements.
At the end of the study, the results are compiled into a formal report.
Required study records and materials are then retained so the work can be reviewed or reconstructed later if necessary.
Together, these controls are intended to protect the integrity and traceability of the resulting safety data.
There is no universal rule that every medical device must complete a GLP study before reaching the market.
The evidence needed for a particular device depends on factors such as its intended use, classification, technological characteristics, patient contact, and identified risks.
Some devices may be supported largely through bench and performance testing.
Others may require biological, animal, or other nonclinical safety studies.
FDA specifically states that GLP applies to nonclinical laboratory safety studies intended to support applications including Investigational Device Exemptions (IDEs) and Premarket Approval (PMA) applications.
For an IDE, if nonclinical laboratory study information is included as part of the report of prior investigations, the sponsor must state whether applicable studies were conducted in compliance with Part 58. If a study was not GLP-compliant, the submission must provide a reason for the noncompliance.
For PMAs, FDA likewise identifies nonclinical laboratory studies as part of the scientific evidence that may support an application.
So the question usually is not:
“Do medical devices require GLP studies?”
A better question is:
“What safety evidence does this device need, and will any of that evidence come from a nonclinical laboratory study that should be conducted under GLP?”
There is no universal GLP protocol for medical devices.
The study should be designed around the specific safety questions raised by the technology.
Depending on the device, nonclinical evaluation might investigate areas such as:
A permanently implanted device, for example, presents different safety questions than a device used temporarily during surgery. A device that delivers energy may introduce different risks than one that functions mechanically.
Those differences can affect the study model, endpoints, duration, controls, and testing strategy.
The key point is that GLP is not itself the safety test.
The safety question comes first. The appropriate study is then designed to answer it under GLP when the regulatory purpose requires that level of control.
It can be tempting to picture device development as a simple sequence:
Prototype → GLP Study → Clinical Trial → FDA Approval
In reality, the process varies considerably between devices.
A simplified pathway might look more like:
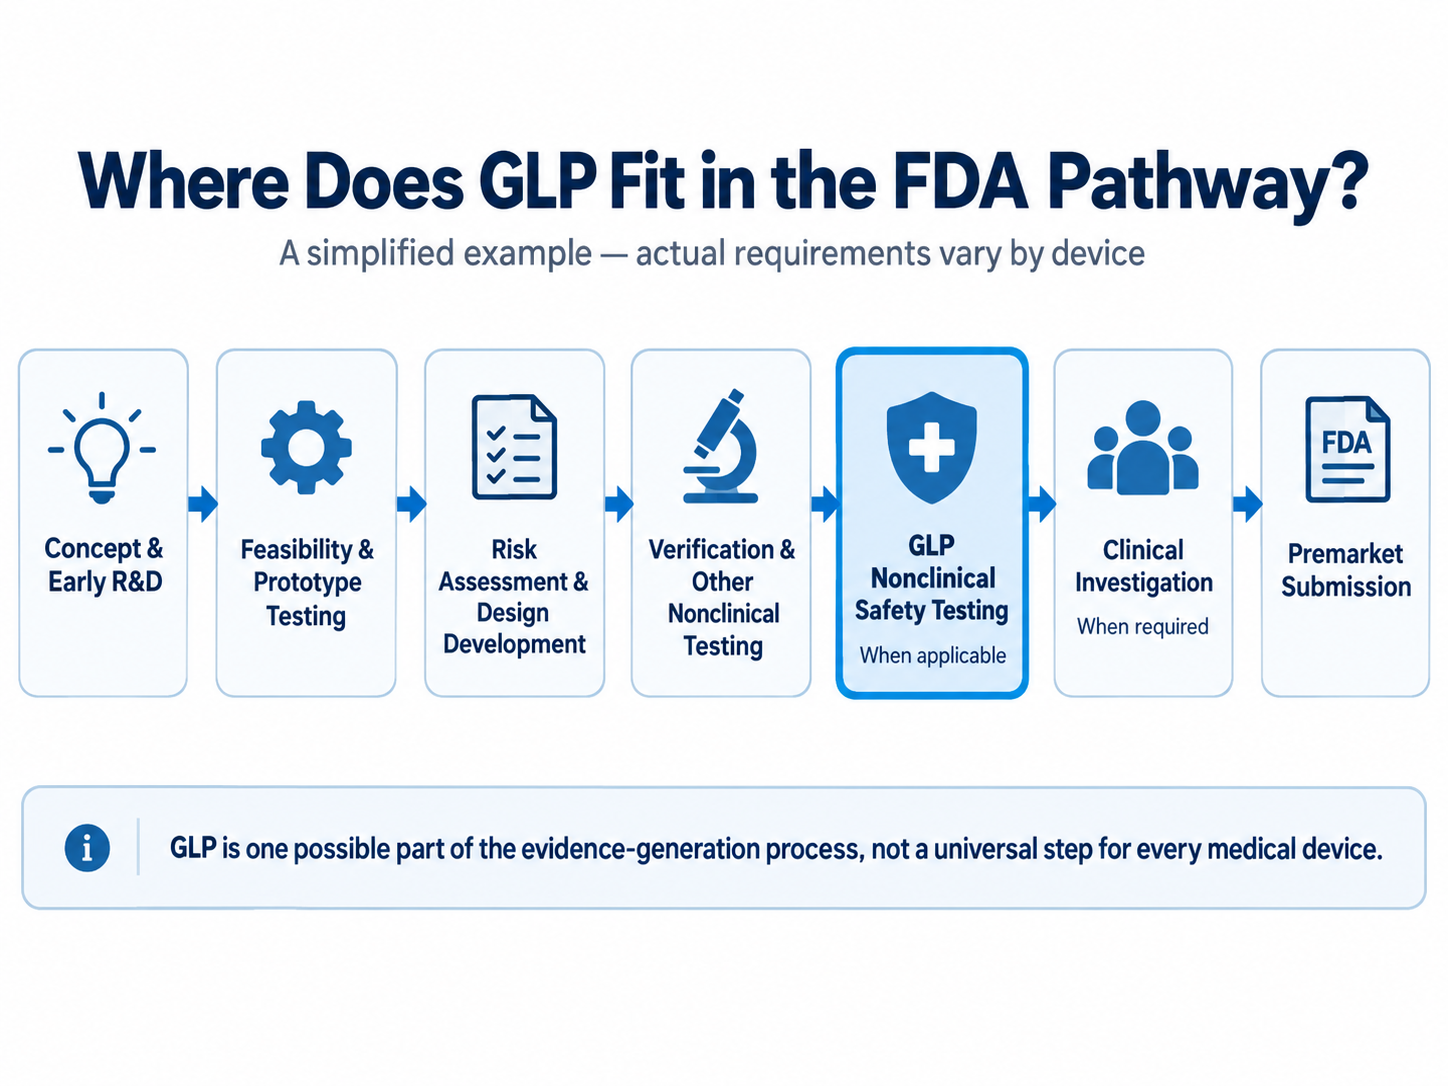
Some activities overlap, and some may occur more than once as the design evolves.
GLP is therefore better understood as one possible part of the evidence-generation process, rather than a universal stage every medical device automatically passes through.
GLP compliance cannot simply be added to a completed experiment after the fact.
The framework affects how the study is conducted from the beginning.
Protocols must be established. Responsibilities must be defined. Equipment and procedures must be controlled. Raw data must be appropriately recorded. Quality assurance oversight must occur. Changes and deviations must be documented.
If a company performs an exploratory study and later decides that the results would be useful in a regulatory submission, that does not automatically make the original study GLP-compliant.
This is why understanding the intended regulatory purpose of a study before it begins is so important.
That does not mean every early experiment should be conducted under GLP. Doing so could add unnecessary structure, time, and cost to testing intended primarily for exploration and development.
Instead, teams need to recognize when they are moving from studies intended primarily for learning into studies intended to become formal regulatory evidence.
The answer is usually device-specific.
Developers can begin by considering:
When the appropriate testing strategy is unclear, early communication with the FDA can also be valuable.
Through the FDA’s Q-Submission Program, companies can request a Pre-Submission, or Pre-Sub, to obtain FDA feedback before committing significant time and resources to a testing strategy.
For example, a developer may ask the FDA whether a proposed nonclinical study is appropriate, whether its endpoints sufficiently address a particular risk, or whether additional testing may be necessary.
A Pre-Sub does not guarantee that the FDA will later accept the resulting data, but it can help identify differences between a company’s proposed strategy and the FDA’s expectations while the study can still be adjusted.
Medical device development involves many regulatory acronyms, and it is easy to confuse some of them.
Applies to certain nonclinical laboratory safety studies and focuses on the quality and integrity of the data generated by those studies.
Addresses clinical investigations involving human participants.
Apply more broadly to how a medical device manufacturer develops, produces, and controls its products and processes.
These systems may interact, but they are not interchangeable.
Operating under a medical device quality system does not automatically make a study GLP-compliant, and conducting a GLP study does not by itself satisfy the broader quality requirements associated with developing a medical device.
GLP can sound complicated when it first appears in a medical device development plan, but the basic concept is relatively straightforward.
Early studies are often about learning:
As development progresses, some studies begin answering a different set of questions:
For applicable nonclinical laboratory safety studies, GLP provides a framework for generating that evidence with the documentation, oversight, and traceability the FDA expects.
That does not mean every medical device needs a GLP study or that every preclinical experiment should be performed under GLP.
It means developers need to understand what question a study is intended to answer, how its data will be used, and what regulatory requirements apply before the study begins.
As with many parts of medical device regulation, addressing those questions early can help keep the testing strategy and regulatory strategy moving in the same direction.
This article is intended as a general educational overview and is not regulatory or legal advice. Requirements vary based on the specific device, intended use, technological characteristics, risks, regulatory pathway, and applicable FDA regulations and guidance.